
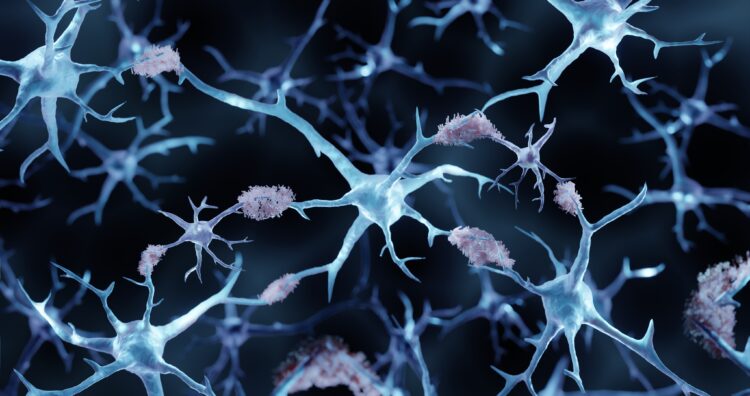
For decades, Alzheimer’s disease research has focused intensely on two primary culprits: beta-amyloid, which forms toxic plaques between brain cells, and tau protein, which creates destructive tangles inside neurons. Yet, a growing consensus within the scientific community suggests that these two biomarkers alone do not fully explain the intricate, devastating progression of the disease.
Now, researchers have uncovered another critical piece of the neurodegenerative puzzle. Under healthy conditions, a protein known as GRK2 plays a vital role in supporting normal neuronal communication and stress responses. However, in the brains of Alzheimer’s patients, this protein can abnormally misfold and clump together. These resulting aggregates directly damage mitochondria—the microscopic energy factories of the cell—triggering a cascade of cellular exhaustion.
A molecular pharmacology team at ETH Zurich analyzed human brain tissue alongside animal models to demonstrate how this altered protein contributes heavily to Alzheimer’s pathology. Led by Professor Ursula Quitterer, a professor of molecular pharmacology at ETH Zurich, the team published their findings in Cell Reports Medicine. In addition to mapping out this pathway, the researchers identified an experimental small-molecule compound capable of blocking the aggregation process in preclinical trials, offering an innovative treatment angle completely distinct from existing therapies.
Why a Vital Protein Turns Destructive
GRK2, or G protein-coupled receptor kinase 2, is an enzyme responsible for regulating how cells interpret and react to external biochemical signals. Active across multiple organ systems, including the heart and the brain, it fundamentally helps cells adapt to stress and maintain structural integrity.
The pathological breakdown begins when GRK2 is altered through normal cellular metabolic processes. The research team discovered that a specific inactivated form, known as serine-670-phosphorylated GRK2 (or phospho-S670-GRK2), accumulates at unusually high levels in the brain tissue of both dementia patients and specialized Alzheimer’s mouse models. When the body fails to properly clear this phosphorylated version of the protein, it rapidly forms sticky, abnormal clusters that prove toxic to neighboring neurons.
Crucially, the study revealed that beta-amyloid and hyperphosphorylated tau—the classic hallmarks of the disease—actively accelerate this misfolding process. This means the toxic elements already known to drive Alzheimer's also destabilize GRK2, establishing a self-perpetuating, vicious loop that worsens neurodegeneration.
Cellular Energy Failures and Mitochondrial Blockades
The core revelation of the study centers on how these GRK2 aggregates directly cripple mitochondrial function. Because neurons require immense amounts of energy to transmit signals and maintain cellular health, any decline in mitochondrial output leaves the brain uniquely vulnerable to permanent damage.
Professor Quitterer’s team discovered that clumped phospho-S670-GRK2 deposits itself directly onto the surface of the mitochondria. Once attached, it interferes with TOMM6 (translocase of outer mitochondrial membrane 6), a crucial outer membrane protein that forms the transport pores regulating the flow of essential substances into and out of the mitochondria. By essentially blocking these vital channels, the aggregates choke off energy production and trigger severe intracellular stress.
This mechanism creates an compounding crisis: beta-amyloid and tau cause GRK2 to clump, the clumped GRK2 starves the cell of energy, and the resulting cellular stress prompts the brain to produce even more beta-amyloid.
A Small-Molecule Candidate Breaks the Loop
Seeking a way to interrupt this destructive feedback mechanism, the research team synthesized several small-molecule chemical formulations and evaluated them in cell cultures and animal models. Among the candidates, an active ingredient designated as "Compound 10" (CPD10) demonstrated remarkable therapeutic efficacy.
Compound 10 works by effectively preventing the inactivated GRK2 proteins from clumping together. Furthermore, it helps restore the protein back to its natural, functional monomer state—the single, unaggregated form required for healthy cell signaling. By keeping the proteins dissolved and monomeric, the molecule protected the mitochondrial pores, reduced cellular stress, and significantly lowered beta-amyloid accumulation.
The preclinical results in live models were highly encouraging. In old-age mice engineered with Alzheimer's features, treatment with Compound 10 noticeably slowed typical neurodegeneration, lowered amyloid deposition, preserved nerve cell function, and prolonged overall survival times. Interestingly, because GRK2 is active body-wide, the treated mice also exhibited secondary anti-aging benefits, including improved heart function and a reduction in age-related graying hair.
Charting a New Frontier for Dementia Research
The underlying research took nearly two decades to complete, primarily due to the complexities of tracking age-related neurological changes over full animal lifespans. Because these findings remain in the preclinical stage, further extensive testing is required to confirm whether the same safety profile and efficacy will translate to human trials.
Currently approved Alzheimer’s drugs primarily focus on mitigating temporary cognitive symptoms or attempting to clear established amyloid plaques, often yielding modest clinical results. The approach pioneered by ETH Zurich is notable because it introduces an entirely separate target. By aiming to regulate intracellular stress and metabolic breakdown rather than simply sweeping away plaques, the study frames Alzheimer’s as a complex, interconnected web of protein aggregation, failed energy systems, and progressive cell death.
The research team has filed a patent application for Compound 10 and is actively seeking corporate bio-pharmaceutical partners to transition the molecule into formal clinical drug development. If successful, this mechanism could one day be utilized alongside existing anti-amyloid therapies, giving clinicians a multi-layered toolkit to attack the disease from multiple biological angles simultaneously.